
Hierarchical polypyrrole-pillared Na2V6O16 architectures enabling high-rate and stable aqueous zinc storage
 0
Graphical Abstract
Abstract
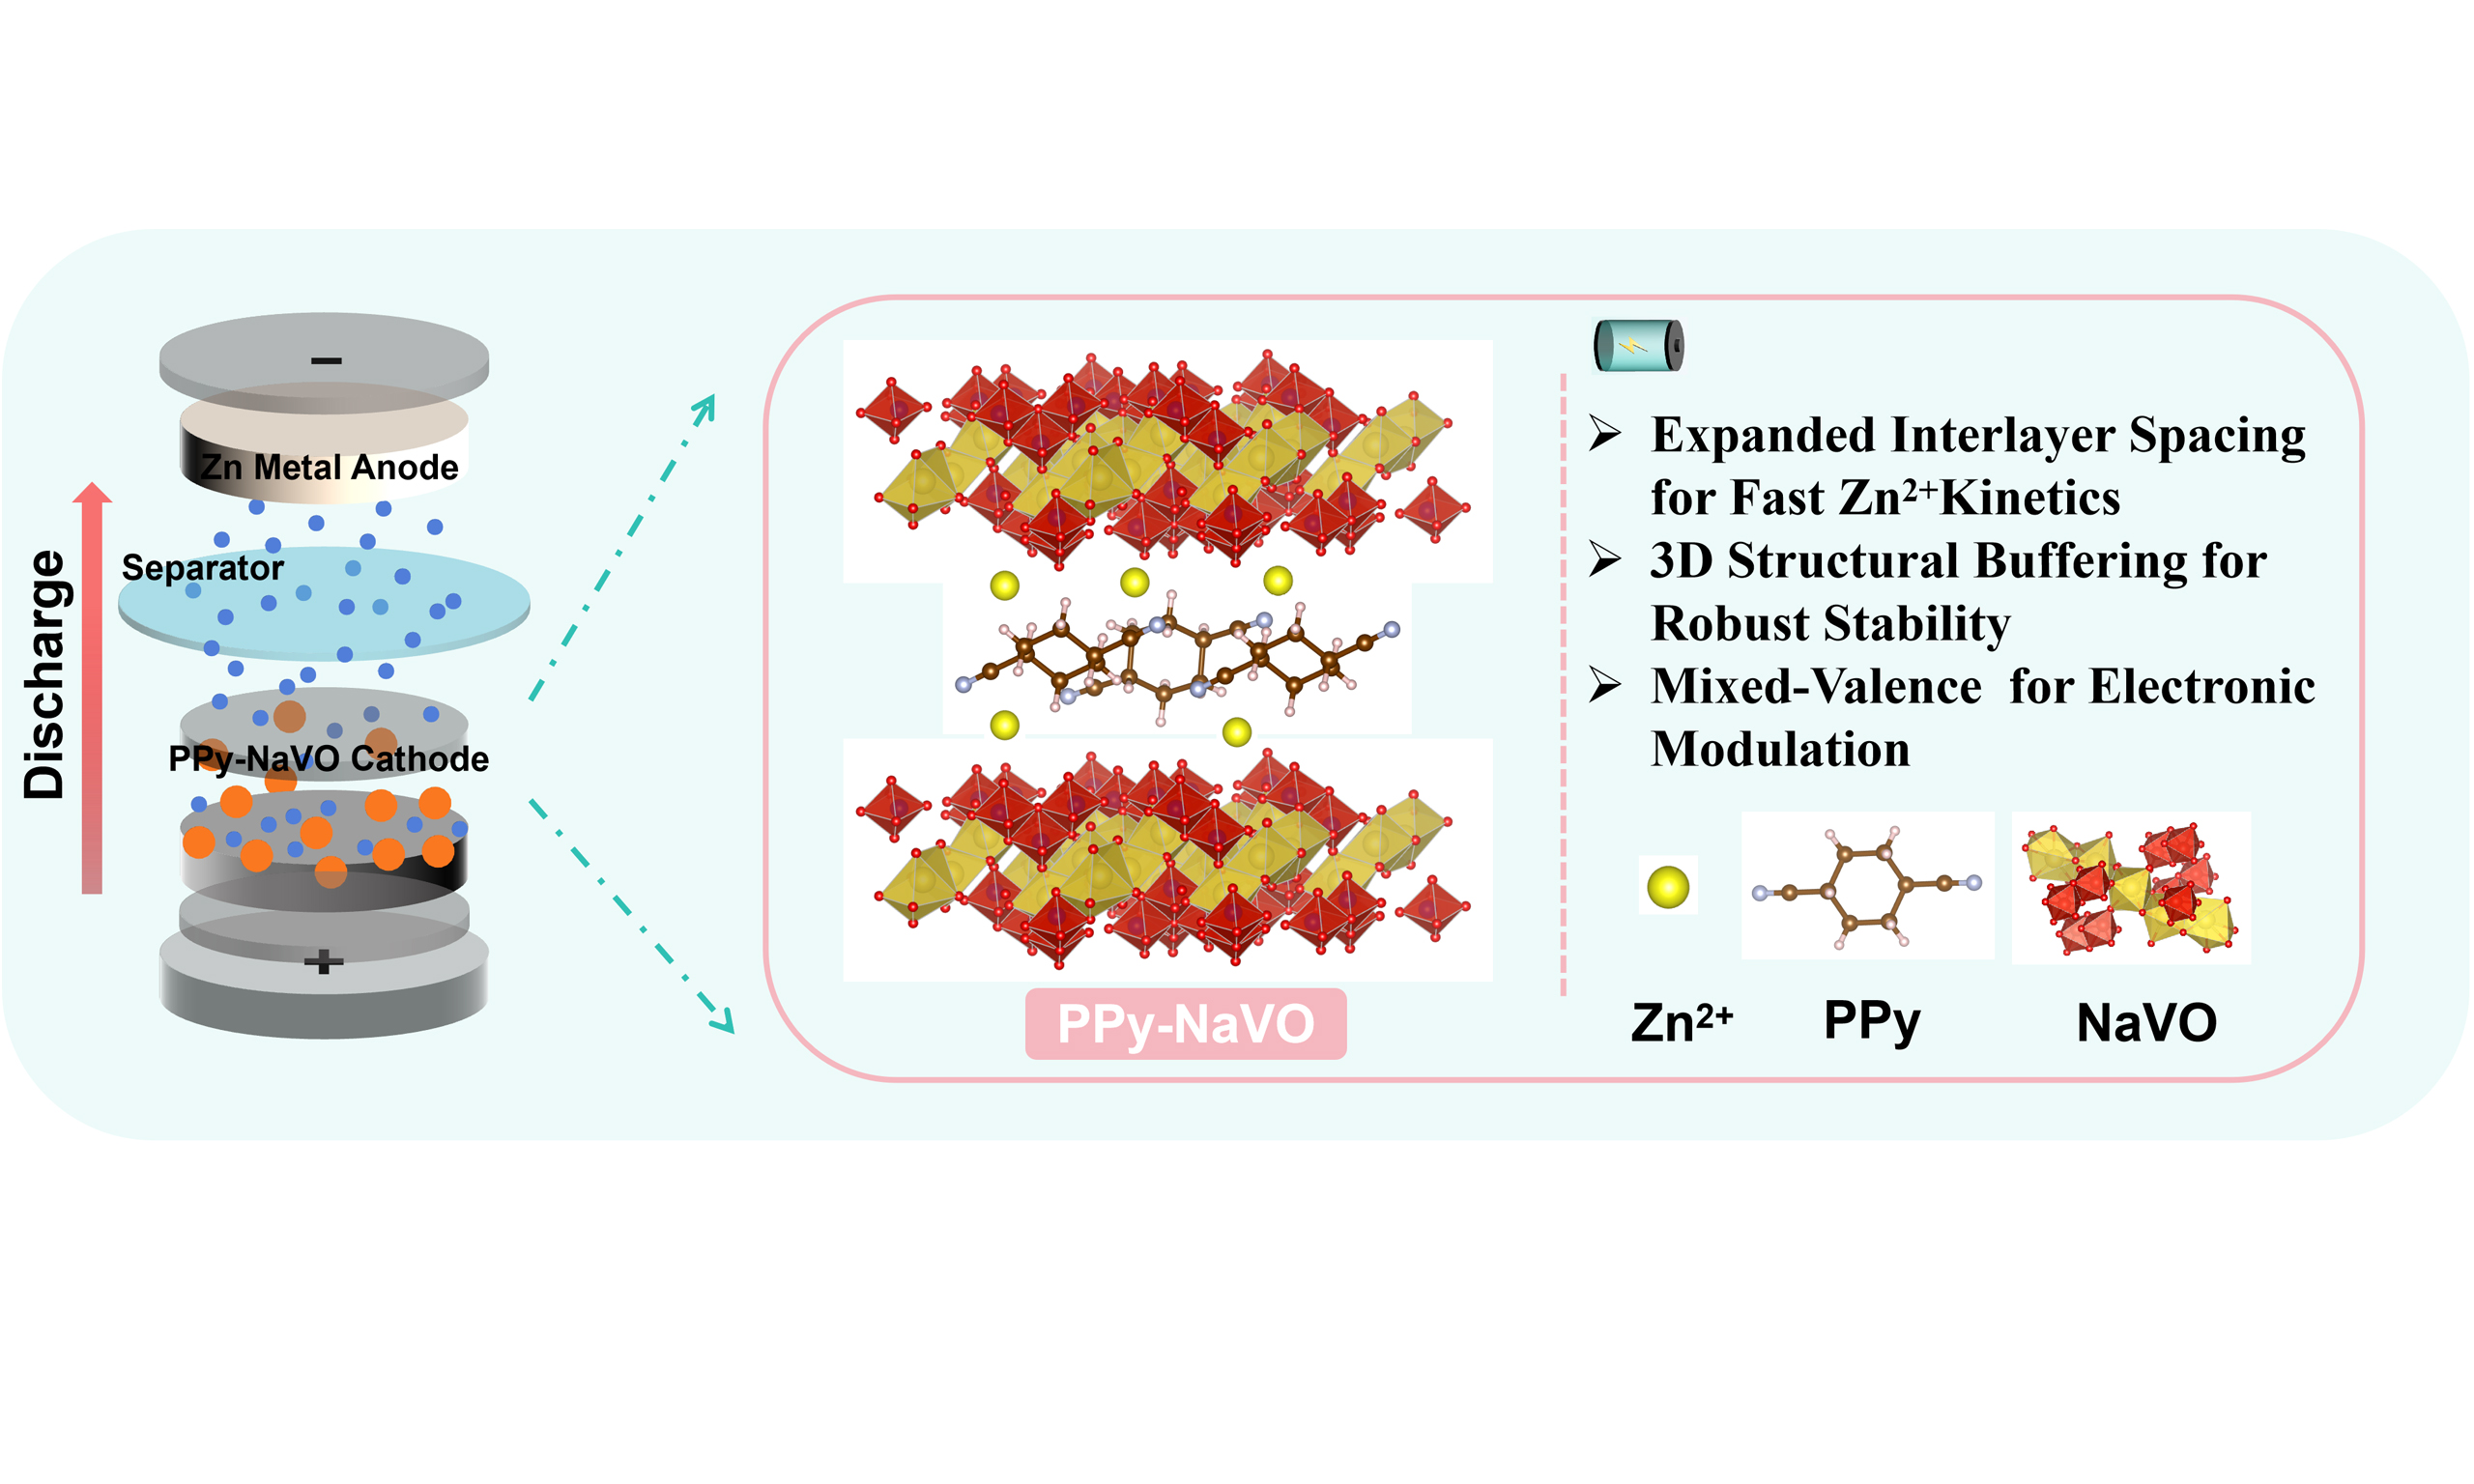
While aqueous Zn-ion batteries represent highly attractive technologies for large-scale grid storage, their practical implementation remains bottlenecked by severe structural degradation and sluggish ion diffusion kinetics in transition-metal oxide cathodes. Herein, a structural buffering hierarchical assembly strategy is proposed to construct a polypyrrole-pillared Na2V6O16 (PPy-NaVO) 3D nanoflower cathode. Structural and morphological analyses reveal that in situ intercalated PPy molecules act as soft pillars to significantly expand the interlayer spacing, which lowers the steric hindrance for ion diffusion and concurrently drives the anisotropic assembly of 1D nanorods into a 3D interconnected porous architecture. This cross-scale structural engineering endows the cathode with intrinsic structural buffering properties to buffer volumetric strain during continuous cycling. Furthermore, the strong interfacial charge transfer between the PPy network and the V-O framework induces a mixed V4+/V5+ valence state. This optimized electronic structure is favorable for interfacial charge transfer and Zn2+ diffusion, minimizing the interfacial resistance and enabling ultrafast Zn2+/H+ co-intercalation kinetics. Consequently, the PPy-NaVO electrode exhibits an impressive reversible capacity
Keywords


 Download PDF
INTRODUCTION
The increasing global requirement for clean energy solutions and grid-scale energy has accelerated the search for post-lithium-ion battery technology that balances safety, cost-effectiveness, and high performance[1-4]. Although the portable electronics sector is dominated by lithium-ion batteries (LIBs), the extensive application of LIBs in large-scale grid storage is severely impeded by both the uneven spatial distribution of lithium reserves and the inherent fire risks associated with combustible organic electrolytes[5-7]. Because of the exceptional theoretical capacity (820 mAh g-1) and inherent safety of metallic zinc anodes, aqueous rechargeable zinc-ion batteries (ARZIBs) have become a very attractive option[8-10]. However, the commercialization of ARZIBs is fundamentally impeded by the lack of ideal cathode materials capable of sustaining rapid and reversible multivalent ion insertion/extraction.
Vanadium-based oxides have garnered a lot of attention from researchers among different potential systems because of their structural variety, tunable coordination environments, and the ability to participate in multi-electron redox reactions, which translate into high theoretical capacities[11,12]. Specifically, layered sodium vanadates, such as Na2V6O16 (NaVO), possess an open two-dimensional framework that is theoretically conducive to Zn2+ diffusion. Nevertheless, the practical performance of NaVO is severely limited by several intrinsic bottlenecks: (i) the strong electrostatic interactions that cause slow diffusion kinetics between the host framework and the high-charge-density Zn2+ ions[13]; (ii) the poor intrinsic electronic conductivity of the vanadium oxide lattice[14]; and (iii) the severe structural instability resulting from periodic lattice strain and active material dissolution during long-term cycling[15]. These issues become even more pronounced under high-rate conditions or mechanical deformation, leading to rapid capacity fading and structural collapse. To overcome these issues, “interlayer engineering” has been extensively investigated as a primary strategy to control the physical and chemical characteristics of layered host materials[16,17]. For example, using Na+/Polyethylene glycol (PEG) co-intercalation, Guo et al. demonstrated an interlayer-engineered ammonium vanadate cathode for rapid and effective Zn-ion storage[18]. Han et al. showed that Ca/F co-doping effectively modifies the layered structure of sodium vanadate, leading to significantly enhanced energy storage performance[19]. However, metal ion pillars often provide limited expansion and may introduce additional electrostatic repulsion, which is insufficient to overcome the high migration barrier of Zn2+[20]. A more advanced strategy in this regard is the intercalation of conducting polymers such as polypyrrole (PPy), poly(3,4-ethylenedioxythiophene) (PEDOT), and polyaniline (PANI)[21-23]. These organic molecules can act as robust molecular pillars to achieve substantial interlayer expansion far beyond that of metal ions, while simultaneously providing continuous pathways for electronic transport and acting as mechanical buffers to accommodate lattice strain. In practice, achieving both fast reaction kinetics and long-term structural stability remains a persistent challenge. Current strategies typically address electronic conductivity or interlayer expansion in isolation, largely overlooking the anisotropic lattice strain and localized stress accumulation during high-rate Zn2+ cycling[24,25]. This oversight inevitably leads to premature structural degradation. Overcoming this limitation requires a hierarchical engineering approach that couples atomic-scale lattice modification with a mesoscale structural buffering architecture.
In this work, a 3D polypyrrole-pillared Na2V6O16 nanoflower architecture (designated as PPy-NaVO) was successfully fabricated via a facile one-step hydrothermal approach for high-rate zinc storage. In this design, the in situ intercalated PPy serves a dual function. Structurally, it acts as molecular pillars to expand the interlayer spacing and directs the self-assembly of 1D nanorods into a 3D structural buffering framework. This hierarchical architecture lowers the steric hindrance for ion diffusion and effectively accommodates anisotropic volume fluctuations. Electronically, strong interfacial charge transfer between the conjugated PPy and the vanadate host induces a V4+/V5+ mixed-valence state, which minimizes the activation energy for electron hopping. Benefiting from the simultaneously optimized ionic and electronic transport, the PPy-NaVO cathode facilitates rapid Zn2+/H+ co-intercalation kinetics, delivering high-rate capability and extended cycling stability. This study offers an effective structural design strategy toward the development of highly stable intercalation cathodes.
EXPERIMENTAL
Materials
All chemical reagents were of analytical grade and were used as received without further purification. Vanadium pentoxide (V2O5), hydrogen peroxide (H2O2, 30%), zinc trifluoromethanesulfonate [Zn(CF3SO3)2], N-methyl-2-pyrrolidone (NMP), and polyvinylidene fluoride (PVDF) were purchased from Shanghai Aladdin Biochemical Technology Co., Ltd. Sodium hydroxide (NaOH) and anhydrous ethanol were obtained from Tianjin Yongda Chemical Reagent Co., Ltd. Hydrochloric acid (HCl) was supplied by Sinopharm Chemical Reagent Co., Ltd. Pyrrole (Py) was purchased from Macklin Reagent. Super P carbon black was obtained from Lion Specialty Chemicals Co., Ltd.
Synthesis of NaVO and PPy-NaVO
First, 45 mL of deionized water was mixed with 1.1 g of V2O5 and 5 mL of 30% H2O2, and the mixture was stirred at 400 rpm for 30 min at 25 °C. After that, 0.32 g of NaOH was added, and stirring was continued at 400 rpm for another 30 min. By adding 0.5 M HCl, the pH of the mixture was adjusted to close to 2. Approximately 40 mL of the prepared yellow suspension was then transferred into a 50 mL Teflon-lined stainless-steel autoclave and heated to 180 °C for 30 h. The dark-red product was collected after cooling to room temperature, repeatedly cleaned with ethanol and deionized water, and then vacuum-dried for 12 h at 60 °C to produce Na2V6O16 (NaVO) nanorods. The PPy·Na2V6O16 (denoted as PPy-NaVO) composite with a 3D mesoporous nanoflower architecture was prepared using a similar procedure, except that 120 μL of Py was introduced into the pale yellow solution and stirred for an additional 30 min before the hydrothermal treatment, ultimately yielding a dark green product.
Characterization
X-ray diffraction (XRD; Bruker D8 AXS, Germany) using Cu Kα radiation (λ = 1.5406 Å) was used to measure the materials’ phase compositions and crystallographic structures. The surface morphologies and elemental distributions were examined using a scanning electron microscope (SEM; Hitachi SU1080, Hitachi, Japan) equipped with an energy-dispersive X-ray spectroscopy (EDS) system. Transmission electron microscopy (TEM; JEM 2100, JEOL, Japan) and high-resolution transmission electron microscopy (HRTEM) were used to examine the microstructures and lattice fringes. The chemical states and valence variations of surface elements were analyzed by X-ray photoelectron spectroscopy (XPS; Thermo Scientific K-Alpha, Thermo Fisher Scientific, USA) using monochromatic Al Kα radiation. Fourier-transform infrared (FTIR) spectra were recorded on a Bruker Tensor II spectrometer (FTIR; Bruker Tensor II, Bruker Corporation, Germany) in the range of 400-4,000 cm-1 using KBr pellets. Raman spectra were collected on a LabRAM HR Evolution Raman spectrometer (Raman; LabRAM HR Evolution, HORIBA, Japan). The specific surface area and pore structure were characterized using an automated surface area and porosity analyzer [Brunauer-Emmett-Teller (BET); Autosorb IQ-Mp, Quantachrome Instruments, USA].
Electrochemical measurements
CR2032 coin cells were assembled for electrochemical testing. The working electrodes were prepared by mixing the active material (NaVO or PPy-NaVO), Super P carbon black, and PVDF binder in NMP at a mass ratio of 7:2:1. The resulting slurry was coated onto a current collector and dried in a vacuum drying oven (DHG-9246A; Shanghai Yiheng Scientific Instrument Co., Ltd.). The mass loading of the active material was approximately 1.5-2.0 mg cm-2, as determined using an electronic analytical balance (Sartorius SQP series; Sartorius Scientific Instruments Co., Ltd.). The cells were assembled using a Zn foil anode, a Whatman glass fiber separator, and a 3 M Zn(CF3SO3)2 aqueous electrolyte, and the assembly was completed with a coin cell crimping machine (MSK-110; Shenzhen Kejing Star Technology Co., Ltd.). All electrochemical measurements were conducted within a voltage window of 0-1.8 V (vs. Zn2+/Zn). Cyclic voltammetry (CV; 0.2-1.0 mV s-1), electrochemical impedance spectroscopy (EIS; 100 mHz-100 kHz, ±5 mV), and galvanostatic intermittent titration technique (GITT) were recorded on a Bio-Logic VMP3 electrochemical workstation (VMP3; Bio-Logic, France). Galvanostatic charge-discharge (GCD) cycling was evaluated using a NEWARE battery testing system (NEWARE; Shenzhen Neware Electronics Co., Ltd.).
RESULTS AND DISCUSSION
The synthesis and structural characteristics of the PPy-NaVO composite are illustrated schematically in Figure 1. Fabricated via a hydrothermal process coupled with in situ polymerization, the composite self-assembles into a unique 3D mesoporous nanoflower architecture. This process enables an effective structural modification of the pristine NaVO host through the bulk intercalation of PPy. The intercalated PPy chains effectively expand the vanadate interlayer spacing, which significantly lowers the migration barrier for solid-state Zn2+ diffusion. Consequently, this structural optimization intrinsically accelerates ionic kinetics, enabling exceptional high-rate zinc storage. SEM and TEM were used to examine the samples’ morphology and microstructure. As seen in Figure 2A and Supplementary Figure 1A and B, the PPy-NaVO composite features a 3D hierarchical nanoflower architecture assembled from interconnected nanorods. In contrast, the pristine NaVO consists of densely aggregated 1D nanorods [Supplementary Figure 1C and D] with corresponding elemental mapping confirming the distribution of Na, V, and O [Supplementary Figure 2]. During synthesis, the in situ polymerized PPy acts as a structure-directing agent through interfacial coordination, driving the anisotropic self-assembly of the nanorods into a 3D mesoporous network. EDS mapping confirms the homogeneous spatial distribution of C, N, Na, V, and O elements throughout the nanoflower structure [Figure 2B-F]. Additionally, the EDS spectra [Supplementary Figure 3] quantitatively verify the substantial incorporation of C and N in the composite, which are absent in the pristine NaVO. The successful incorporation of PPy into the vanadate framework is consistent with a uniform distribution of C and N. The pristine NaVO exhibits lattice fringes with a d-spacing of 1.542 Å, corresponding to the (431) plane [Figure 2G and H]. For PPy-NaVO, lattice fringes with a d-spacing of 1.804 Å are observed, corresponding to the (411) plane [Figure 2I and J]. These observations indicate that PPy incorporation alters the local lattice structure of the vanadate framework. Furthermore, the interconnected 3D architecture and intrinsic mesoporosity act as mechanical buffers to accommodate localized stress. This configuration accommodates the anisotropic volume changes during continuous Zn2+ (de)intercalation, consequently maintaining the electrode's structural stability throughout extended cycling.
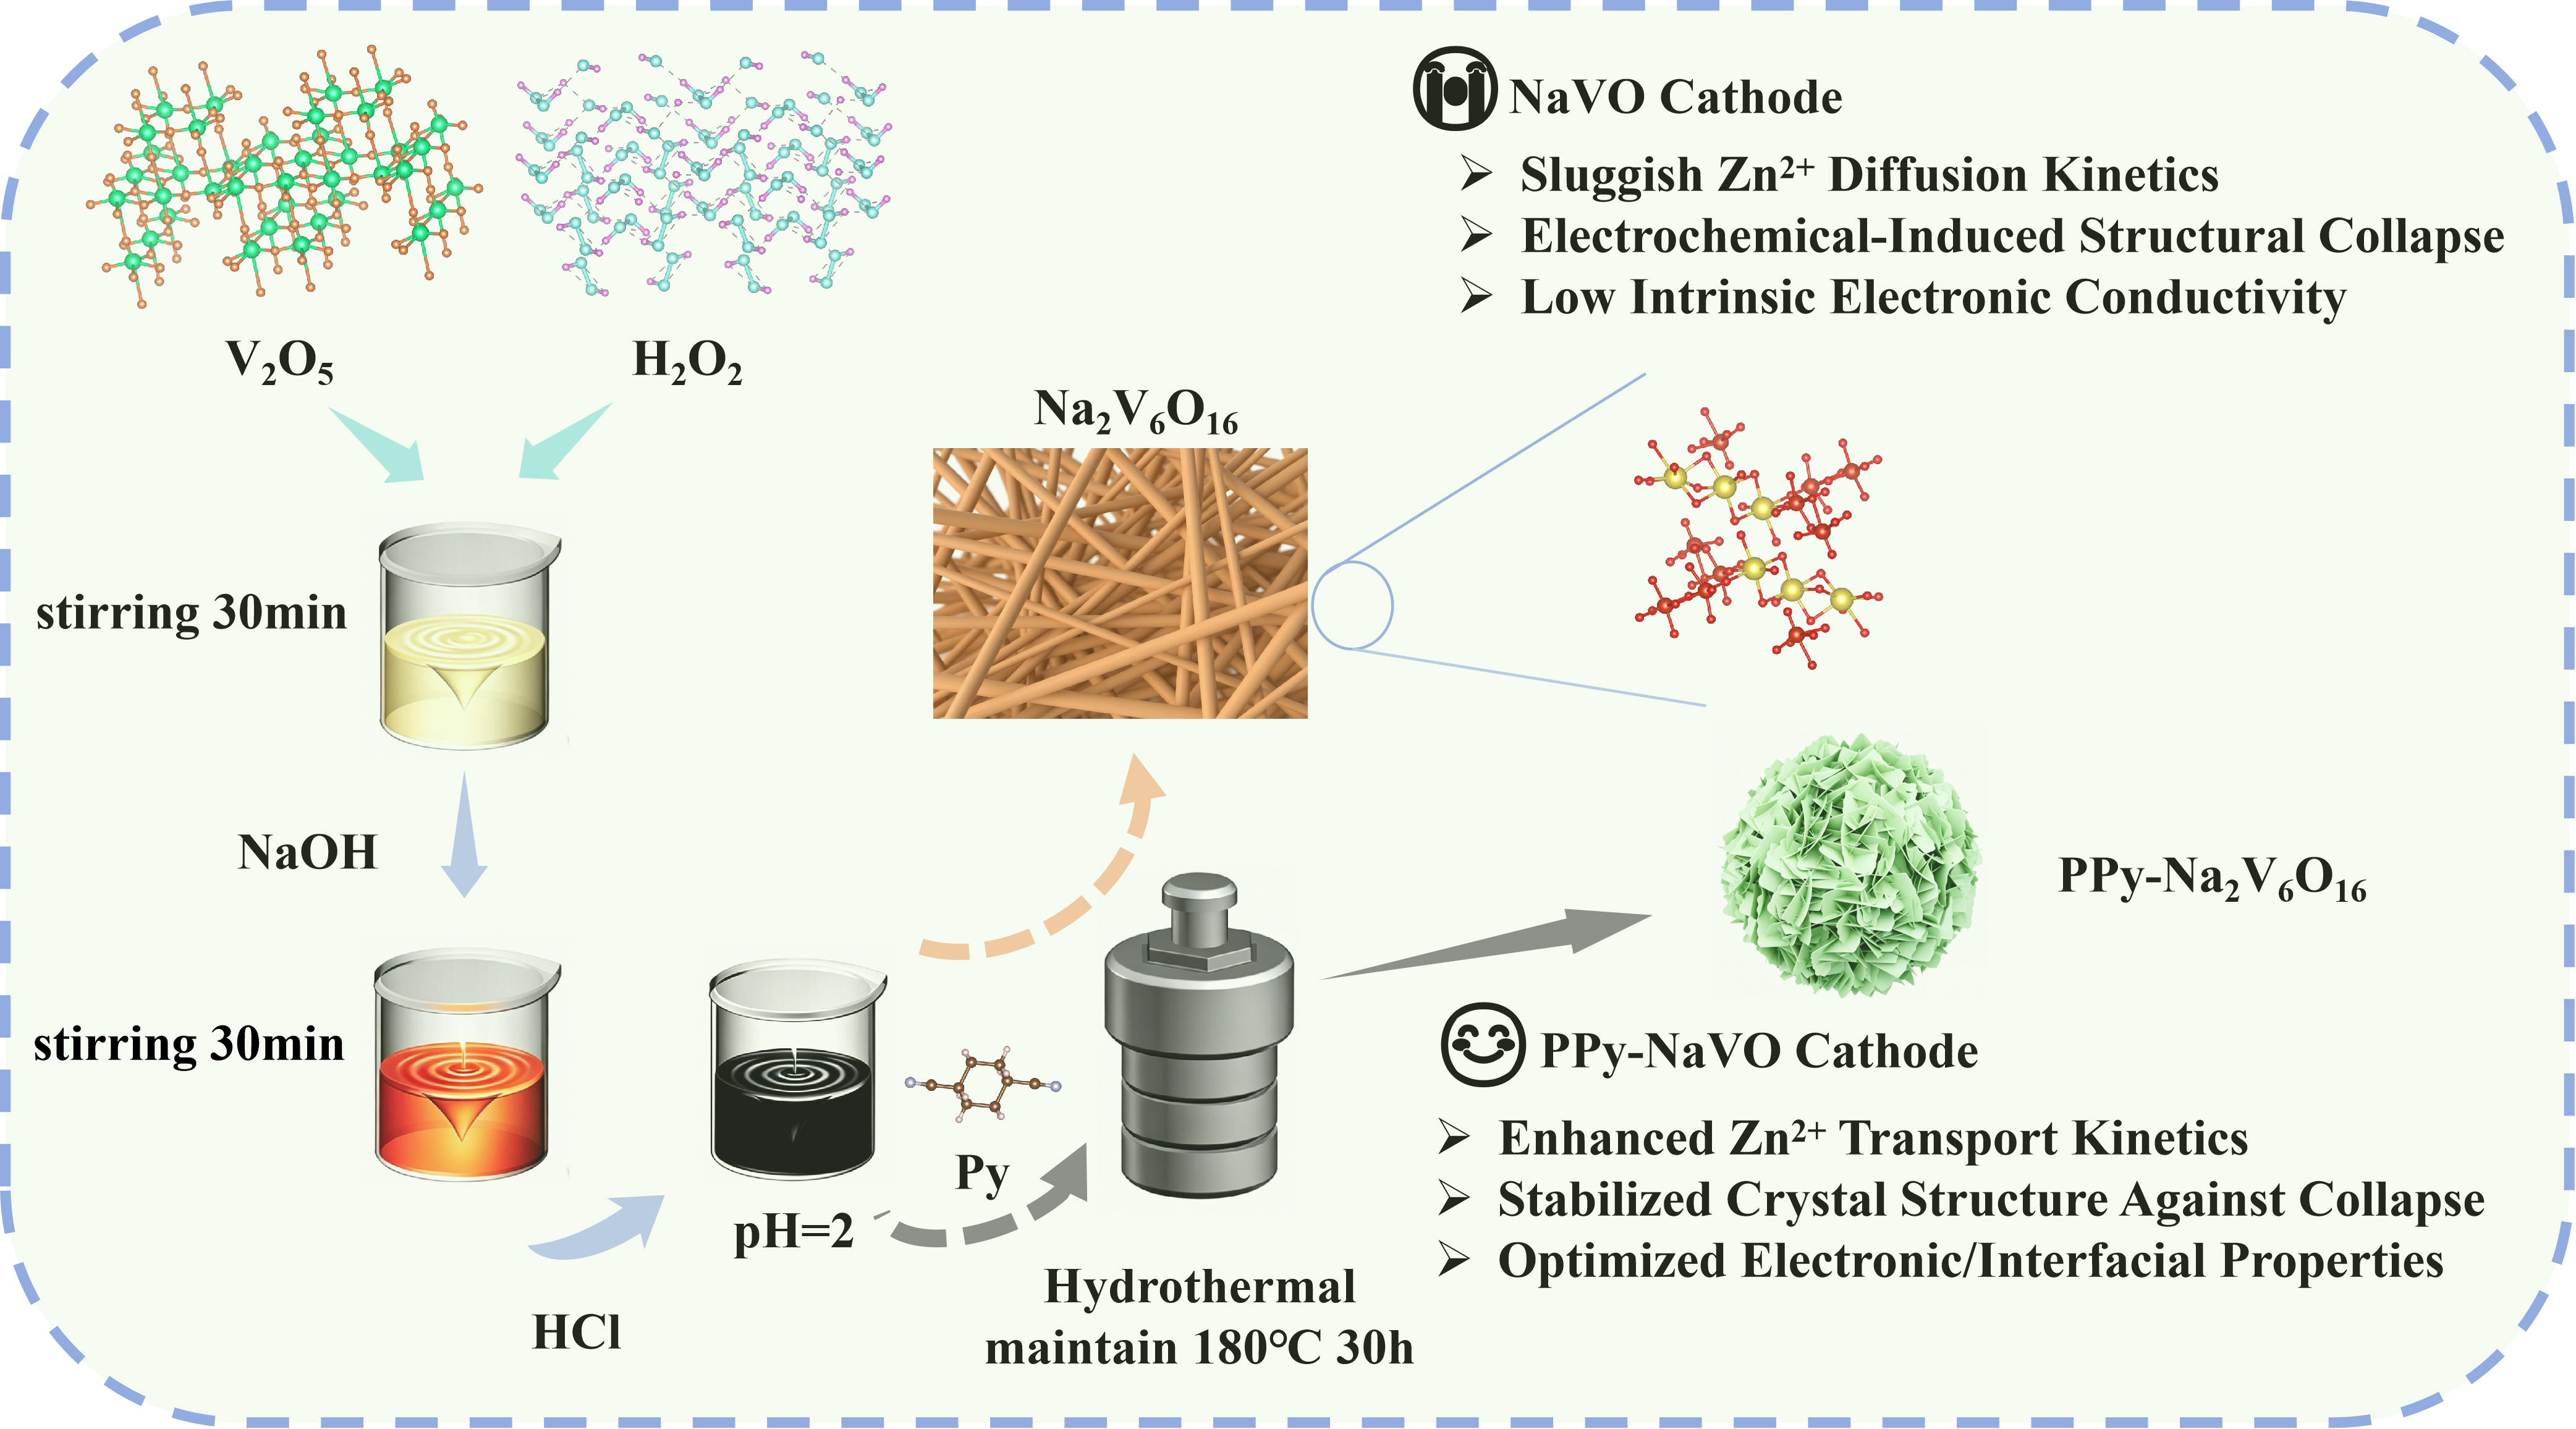
Figure 1. Schematic diagram of the synthesis pathway for the PPy-NaVO composite. PPy-NaVO: Polypyrrole-pillared Na2V6O16.
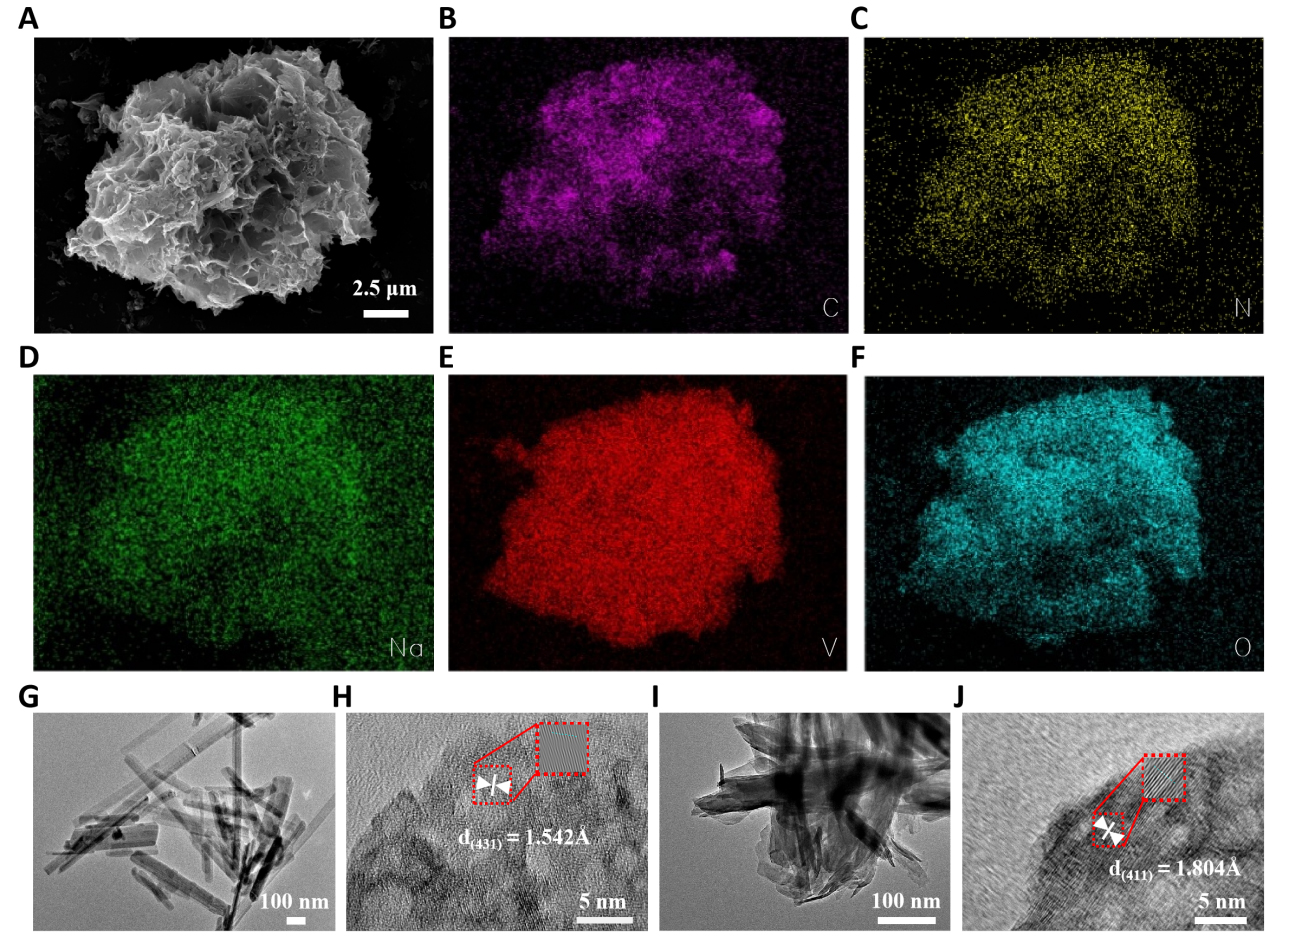
Figure 2. (A) SEM image of PPy-NaVO; (B-F) EDS mapping images of (A); (G) TEM image of NaVO; (H) HRTEM image of NaVO; (I) TEM image of PPy-NaVO; (J) HRTEM image of PPy-NaVO. EDS: Energy-dispersive X-ray spectroscopy; TEM: transmission electron microscopy; HRTEM: high-resolution transmission electron microscopy; SEM: scanning electron microscope; PPy-NaVO: polypyrrole-pillared Na2V6O16.
The crystalline structures and phase compositions were further determined by XRD. The pure layered phase (PDF#22-1412) has been identified from the pristine NaVO diffraction pattern [Figure 3A]. After PPy intercalation, the characteristic (001) diffraction peak shifts from 11.2° to 9.88° [Figure 3B]. This shift corresponds to an increase in the interlayer distance from 0.79 nm to 0.89 nm, according to Bragg’s equation. This expanded interlayer spacing lowers the migration barrier for solid-state Zn2+ diffusion and provides additional free volume to accommodate the intercalated ions and volume changes during cycling. The chemical composition was examined using FTIR spectroscopy [Figure 3C]. The low-wavenumber V-O vibrations in PPy-NaVO are preserved, indicating that the basic vanadate framework is maintained after intercalation. Additionally, other absorption bands appear at 1,410 and 765 cm-1, which are attributed to the pyrrole ring’s C-N stretching and C-H out-of-plane bending vibrations, respectively[26]. The broad band near 1,600 cm-1 is attributed to the overlapping signals of the intercalated water bending vibration and the PPy C=C skeletal stretching[27]. To elucidate the interfacial electronic structure and chemical states of the composite, XPS was employed. Na, V, O, C, and N are present in the survey spectrum [Supplementary Figure 4], with no identified impurities. The high-resolution C 1s spectrum of PPy-NaVO [Figure 3D] displays three peaks at 284.7, 285.8, and 287.9 eV, which are assigned to C-C, C-N, and C-O bonds, respectively[28]. Three peaks can be seen in the N 1s spectra [Figure 3E] at 399.7, 400.7, and 402.1 eV, which correspond to the pyrrole ring’s -NH-, -N+H-, and =N+- species[29]. The existence of positively charged nitrogen species suggests that the intercalated PPy is in an oxidatively doped state, which enhances the composite’s electrical conductivity. The V 2p high-resolution spectra [Figure 3F] further verify the interfacial charge transfer. Pristine NaVO exhibits a typical spin-orbit doublet at 517.3 eV (V 2p3/2) and
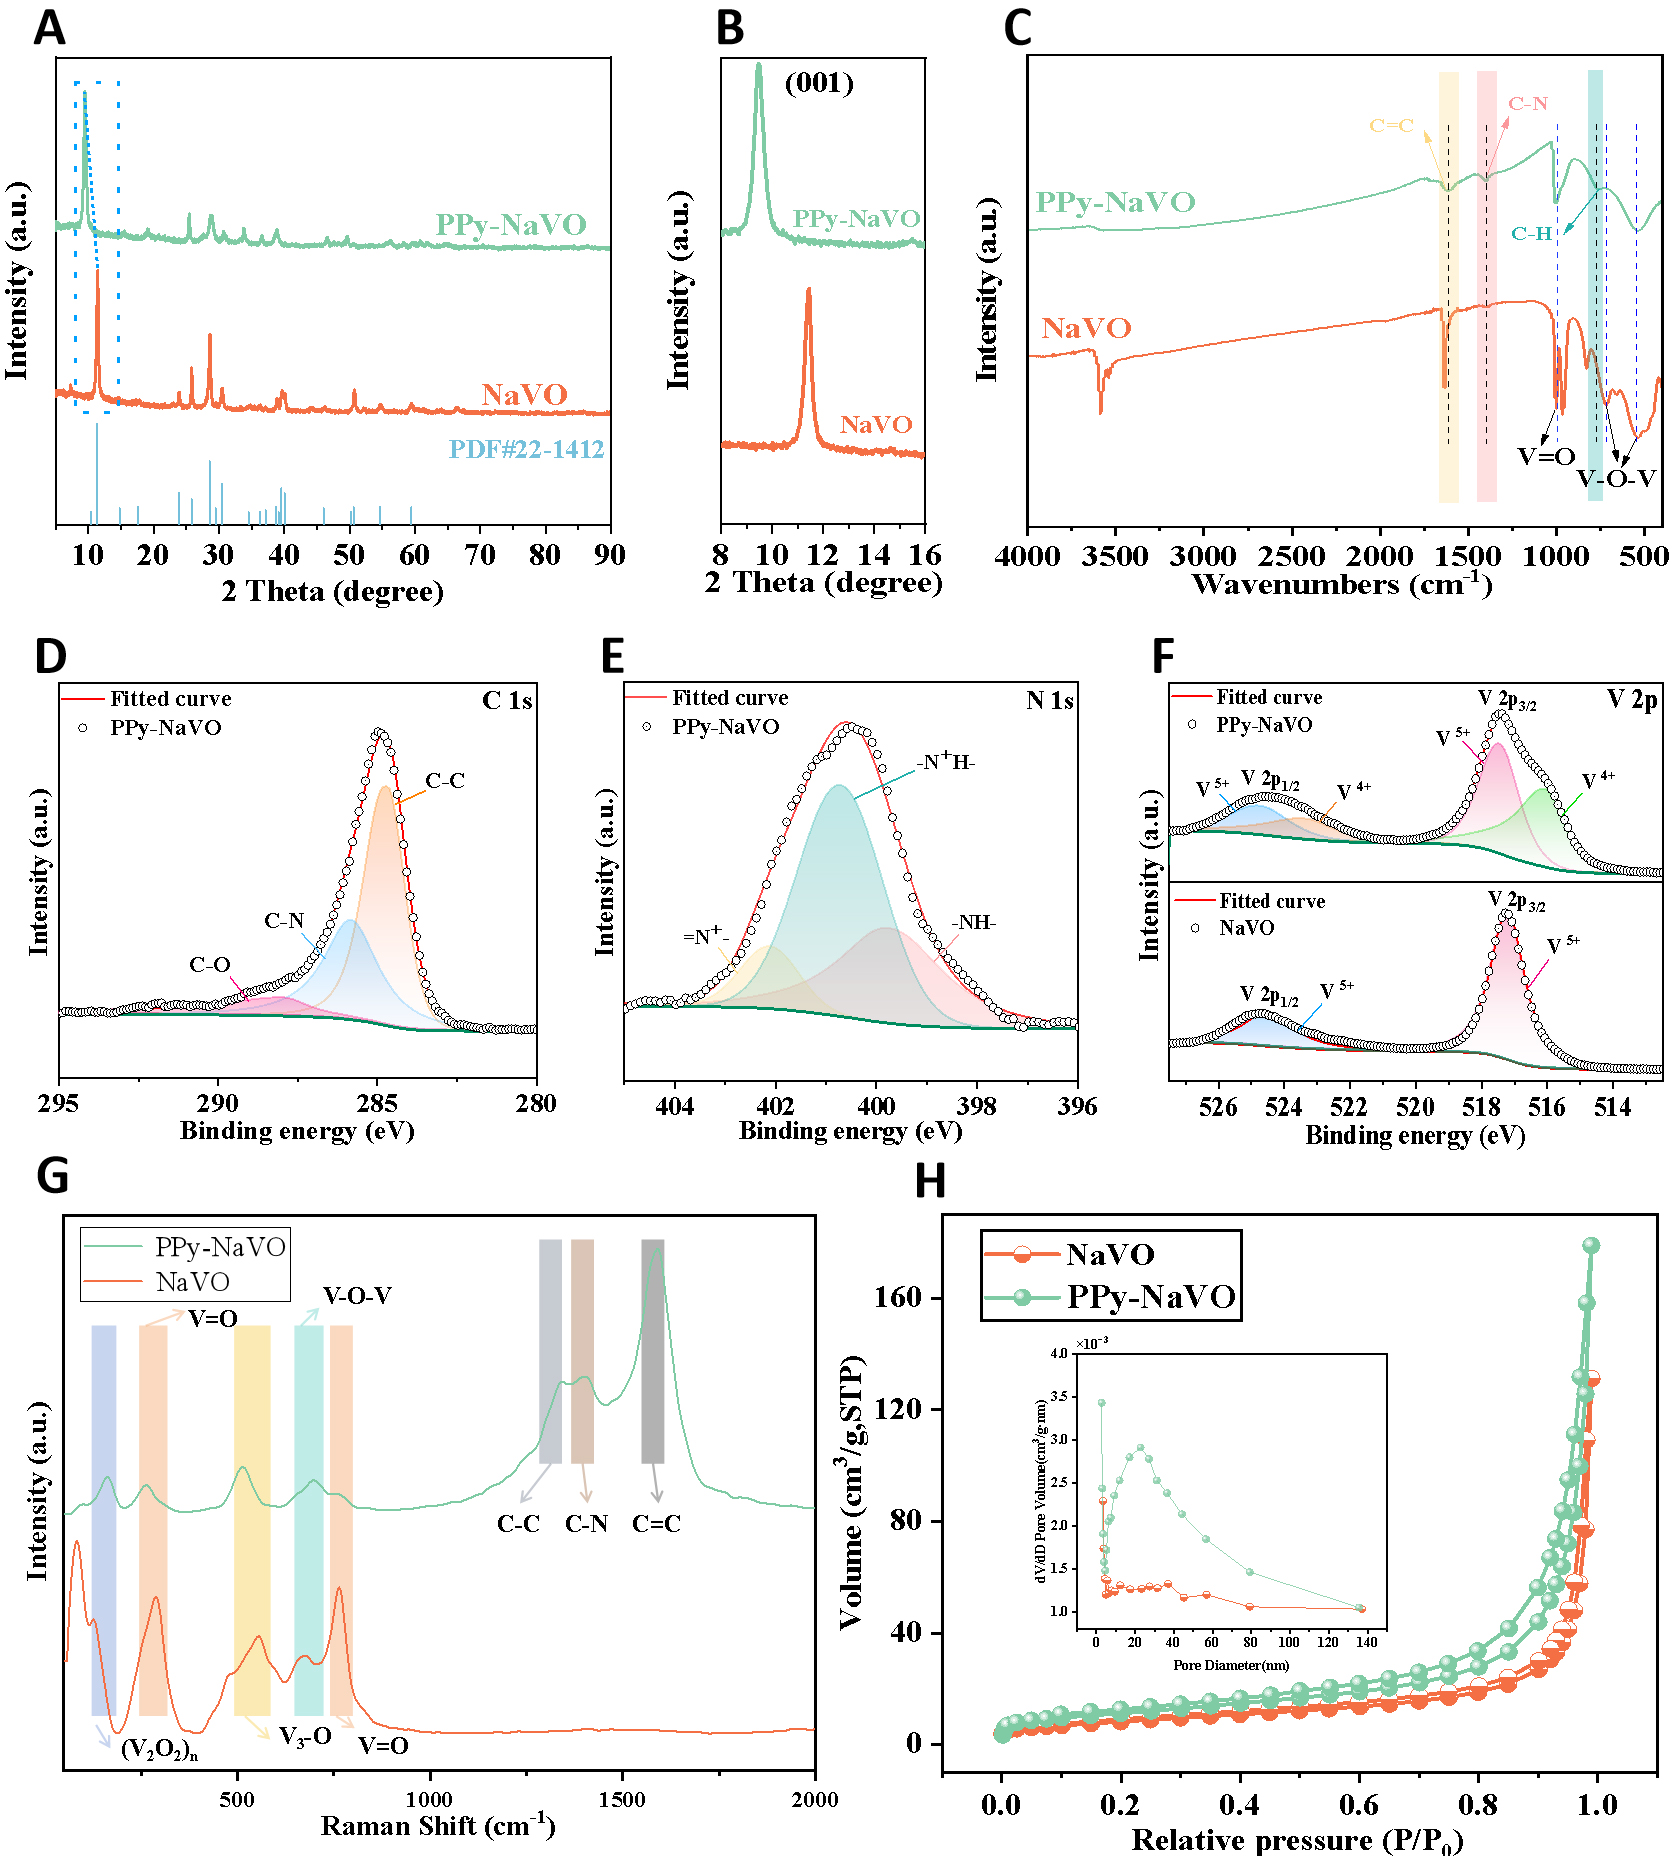
Figure 3. Chemical structure characterization of PPy-NaVO. (A) Full XRD patterns; (B) Enlarged view of the (001) diffraction peak; (C) FTIR spectra; (D-F) High-resolution XPS spectra; (G) Raman spectra; (H) Pore size distribution curves (inset) and N2 adsorption-desorption isotherm curves. XRD: X-ray diffraction; XPS: X-ray photoelectron spectroscopy; FTIR: Fourier transform infrared spectroscopy; PPy-NaVO: polypyrrole-pillared Na2V6O16.
To evaluate the electrochemical zinc storage potential, CV was performed, and the profiles obtained at
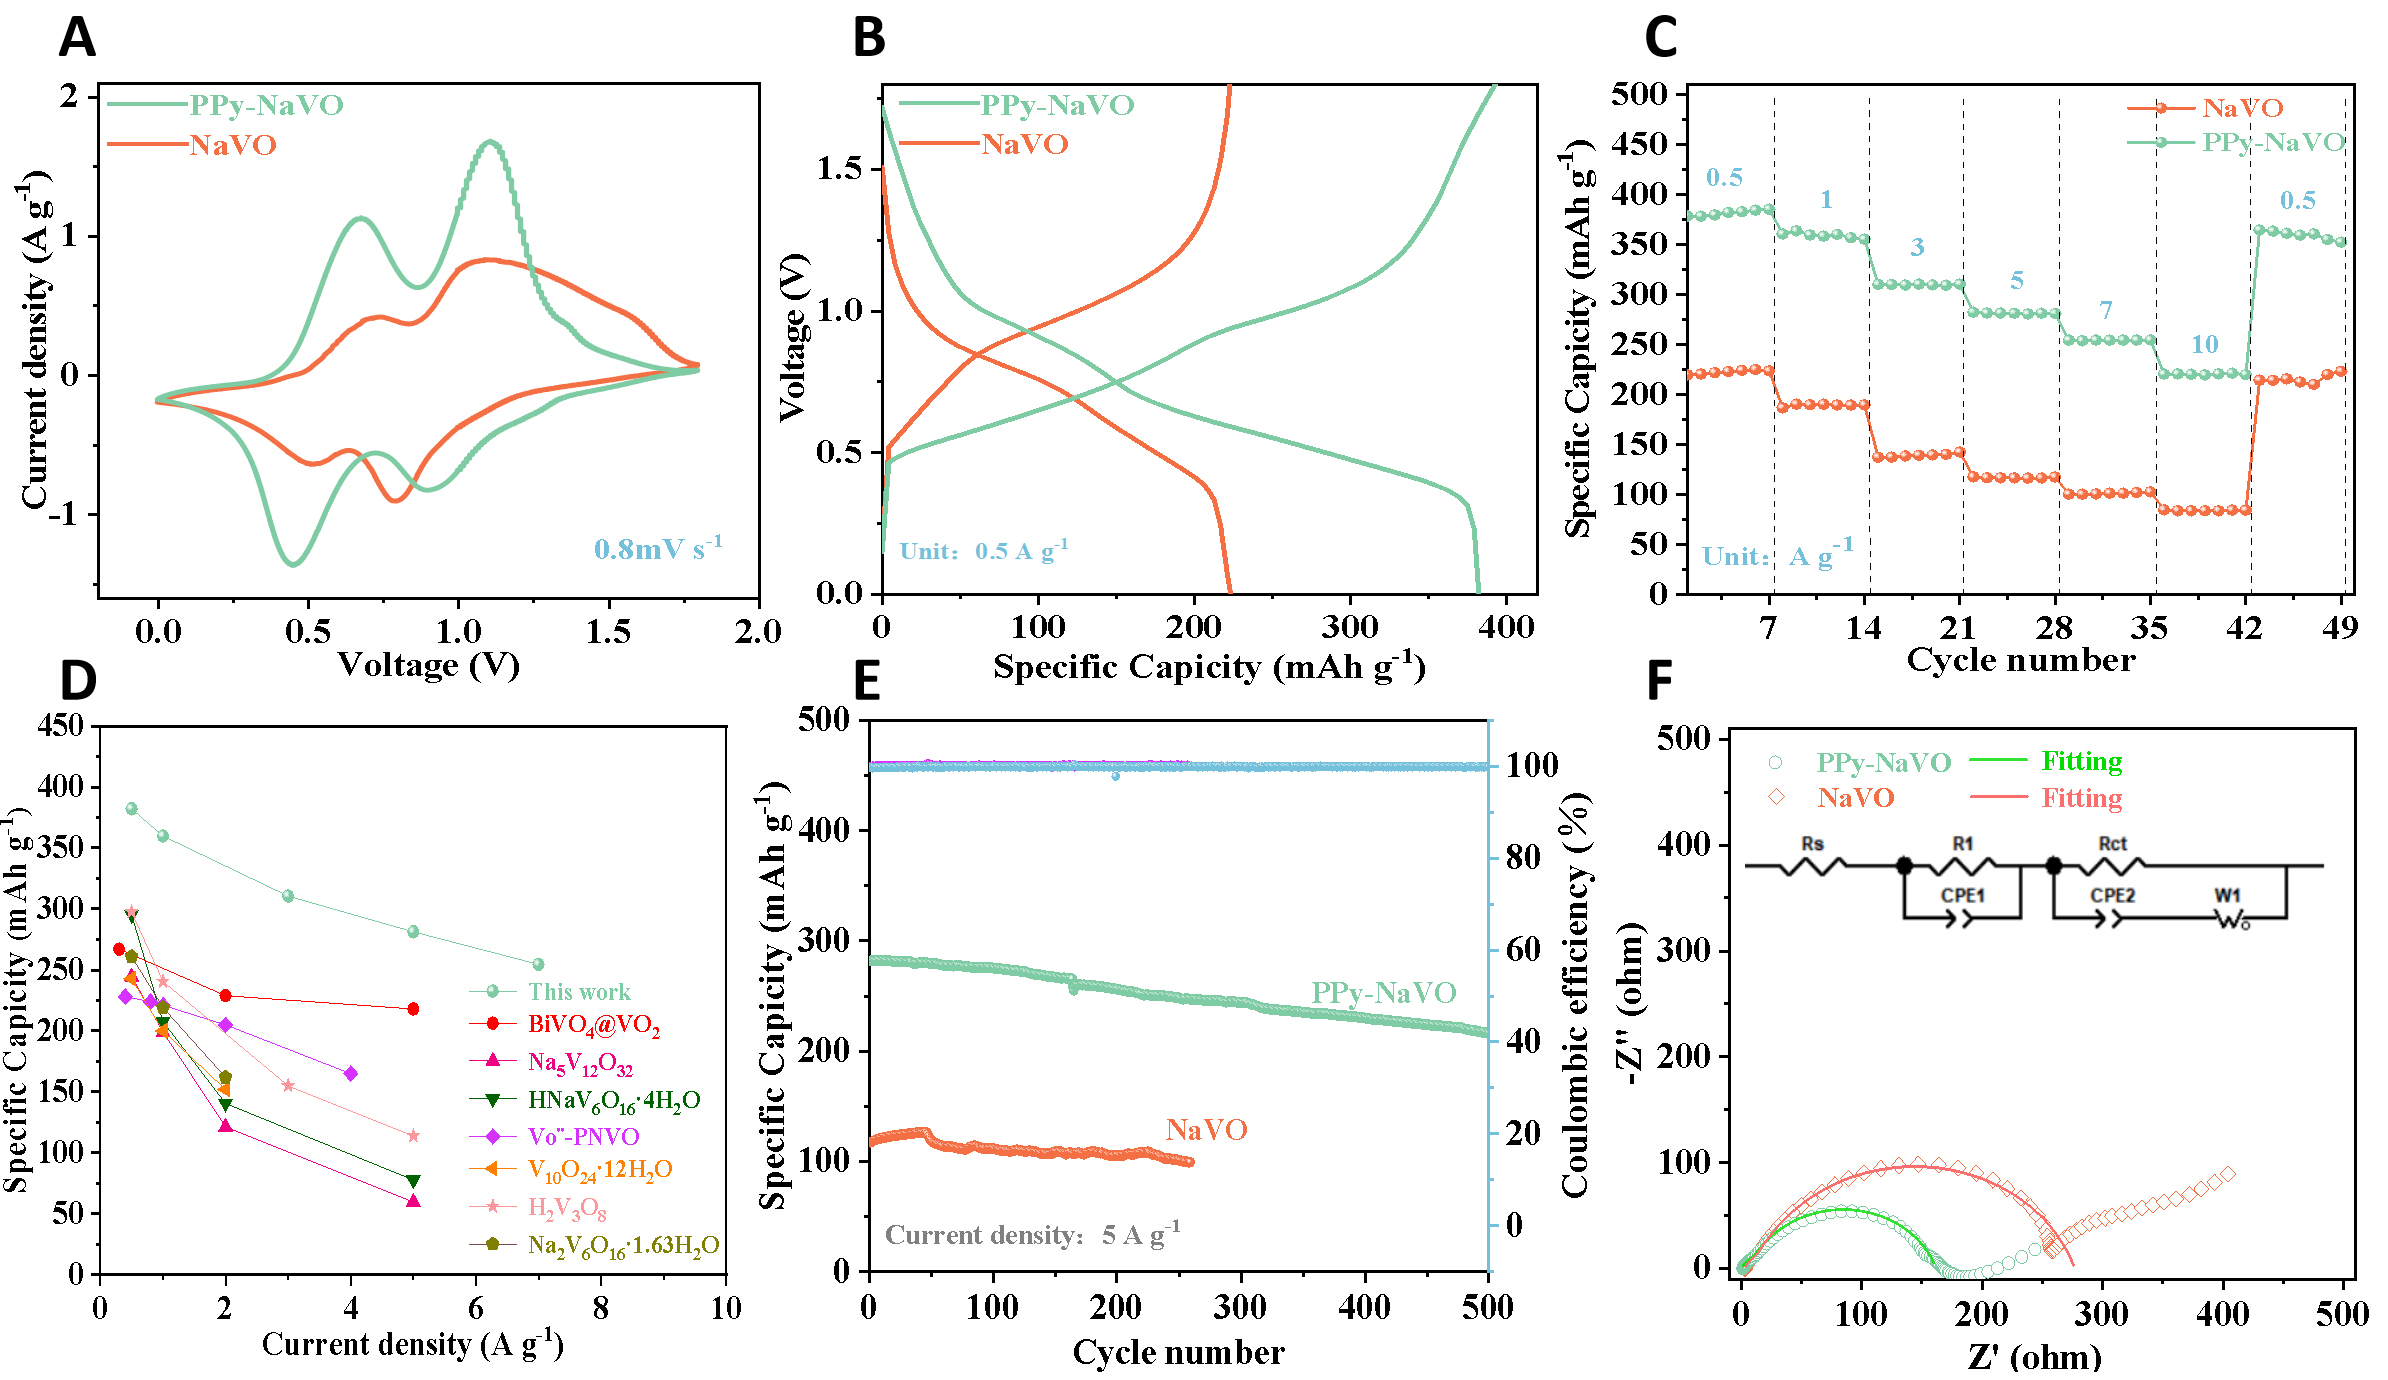
Figure 4. (A) CV curves; (B) GCD curves; (C) Rate performance; (D) Capacity comparison with reported vanadium-based AZIB cathodes; (E) Cycling stability at 5 A g-1; (F) Nyquist plots (EIS). CV: Cyclic voltammetry; GCD: galvanostatic charge-discharge; EIS: electrochemical impedance spectroscopy; AZIB: aqueous zinc-ion battery; PPy-NaVO: polypyrrole-pillared Na2V6O16.
The Zn2+ storage kinetics were investigated using CV at different scan rates [Figure 5A]. The peak shapes are retained at all scan rates, indicating reversible redox processes. The four redox peaks of PPy-NaVO have b values of 0.88, 0.64, 0.78, and 0.71 when the peak current (i) versus scan rate (v) is analyzed using i = avb [Figure 5B]. These b values exceed those of the pristine NaVO [Supplementary Figures 9 and 10], indicating a charge storage mechanism comprising both diffusion-controlled intercalation and surface-controlled capacitive processes. The capacitive contribution of PPy-NaVO [Figure 5C] increases from 53.86% at 0.2 mV s-1 to 71.61% at 1.0 mV s-1. For pristine NaVO, the capacitive ratio ranges from 46.41% to 66.37% at the same scan rates [Supplementary Figures 11 and 12]. As PPy-NaVO has more sites for surface-controlled charge transfer owing to its larger specific surface area, it has a more significant capacitive contribution. GITT was used to calculate solid-state Zn2+ diffusion coefficients [Figure 5D]. The V4+/V5+ mixed-valence state in PPy-NaVO effectively lowers the Zn2+ transport barrier, ensuring robust DZn values [Figure 5E]. The PPy-NaVO//Zn cell exhibits an energy density of 278.99 Wh kg-1 with a power density of 365.07 W kg-1, illustrated in the Ragone plot [Figure 5F]. To demonstrate practical applicability, a series connection of two PPy-NaVO//Zn coin cells was utilized to power an electric fan [Supplementary Figure 13], while three cells connected in series successfully illuminated an Light-Emitting Diode (LED) display [Supplementary Figure 14]. The structural evolution and charge storage mechanism of the PPy-NaVO cathode were investigated using ex-situ characterizations [Figure 6A]. During the discharge process, the (001) diffraction peak shifts to a smaller angle in the ex-situ XRD patterns [Figure 6B and C]. This shift corresponds to the increase in interlayer space due to hydrated Zn2+ intercalation. The (001) peak recovers to the initial state after charging to 1.15 V, indicating reversible lattice variation. Additionally, new diffraction peaks emerge during discharge, corresponding to Zn3(OH)2V2O7·2H2O (ZVO) and a basic zinc salt (BZS)[37,38]. Upon fully charging the electrode, the intensity of these diffraction peaks decreases significantly, although weak residual signals remain. This highly reversible precipitation-dissolution behavior suggests a co-insertion mechanism[39]. The chemical state evolution was examined using ex-situ XPS. The high-resolution Zn 2p spectrum [Figure 6D] reveals no initial zinc signal. After discharging to 0 V, the Zn 2p doublet appears at 1,023.0 eV and 1,046.2 eV, confirming Zn2+ intercalation. Following the charge process, the Zn 2p signal intensity decreases, indicating Zn2+ extraction. The weak residual signal is attributed to trapped Zn2+ or residual precipitates[40]. The V 2p3/2 spectrum [Figure 6E] tracks the vanadium valence states. The pristine PPy-NaVO electrode presents a V4+/V5+ mixed-valence state. A V3+ peak emerges, and the V5+ component decreases during discharge, reflecting a steady reduction via V5+ to V4+ and then to V3+. Upon full charging, the V 2p3/2 spectrum returns to the initial mixed-valence state[41]. The PPy-NaVO cathode’s electrochemical mechanism is shown in Figure 6F. In addition to structural expansion/contraction and multielectron vanadium redox reactions, the process includes a reversible Zn2+/H+ co-intercalation. The intercalated PPy network and the mesoporous architecture provide continuous electron and ion transport pathways. Furthermore, this structural configuration acts as a mechanical buffer to accommodate localized stress caused by lattice volume changes, keeping the structural stability of the cathode during repeated cycling.
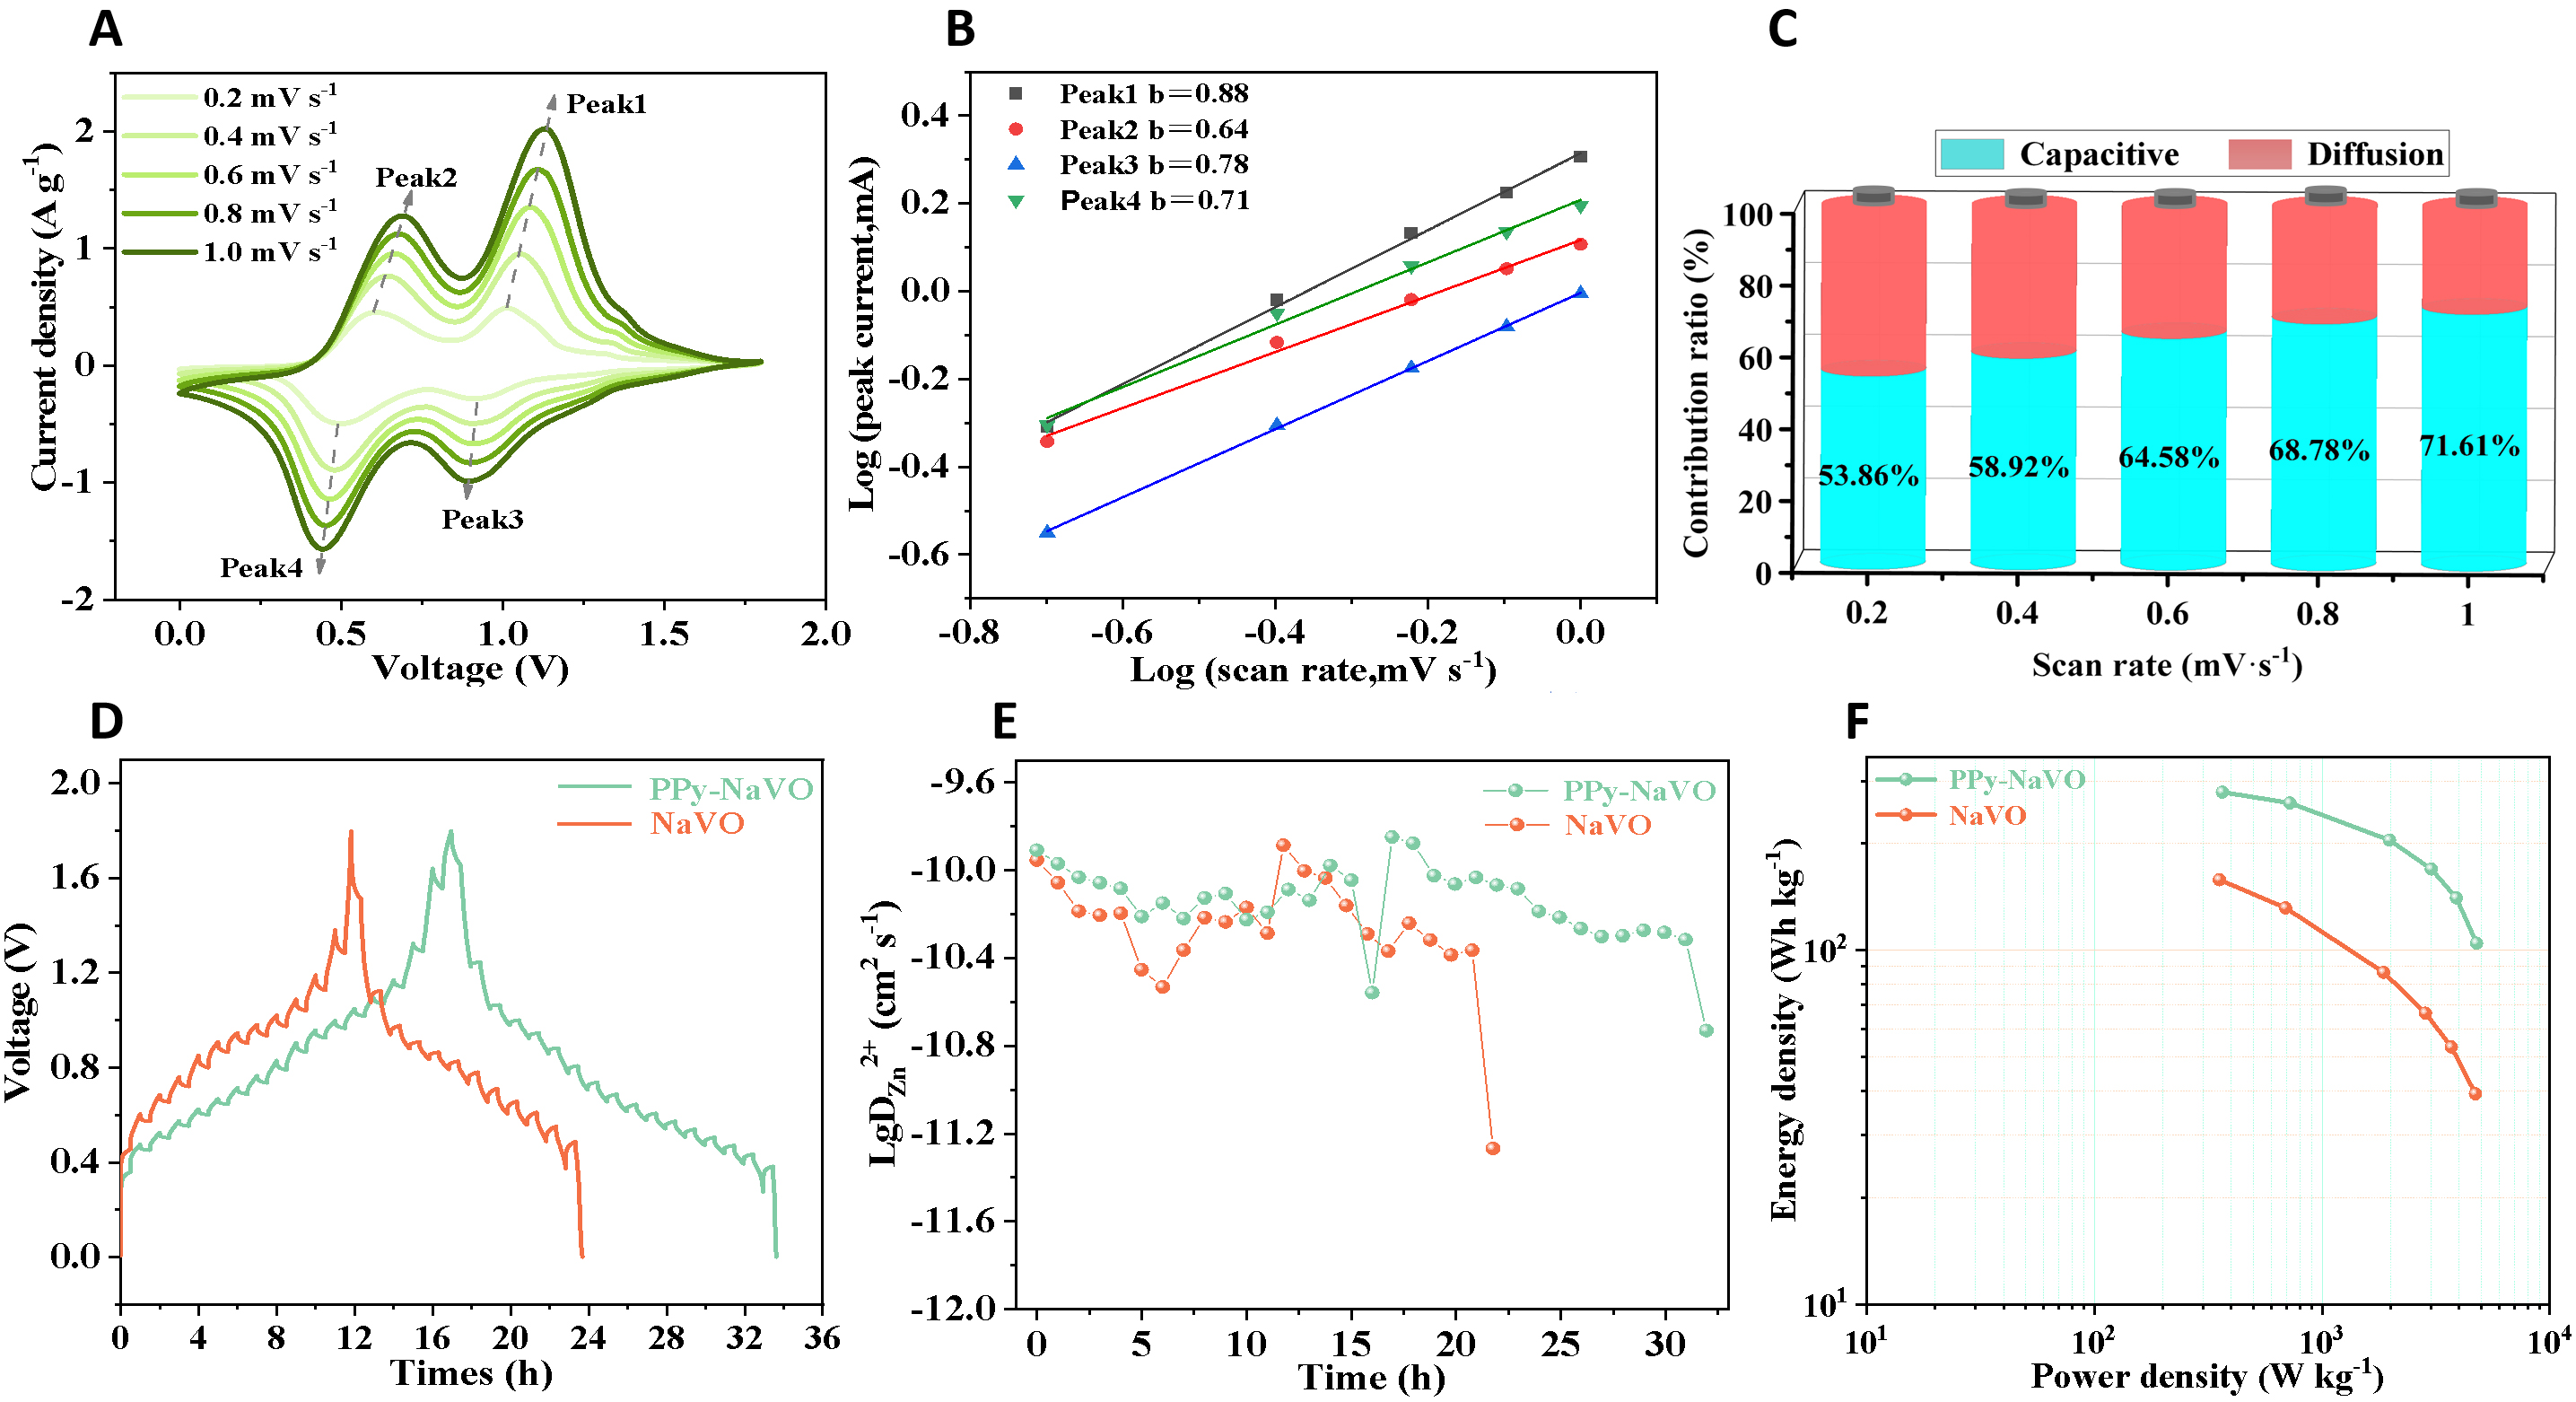
Figure 5. (A) CV curves; (B) Log(i) versus log(v) fitting plot; (C) Capacitive contribution of PPy-NaVO at different scan rates; (D) GITT charge-discharge comparison curves; (E) Comparison of Zn2+ diffusion coefficients; (F) Ragone plot of NaVO and PPy-NaVO. CV: Cyclic voltammetry; GITT: galvanostatic intermittent titration technique; PPy-NaVO: polypyrrole-pillared Na2V6O16.
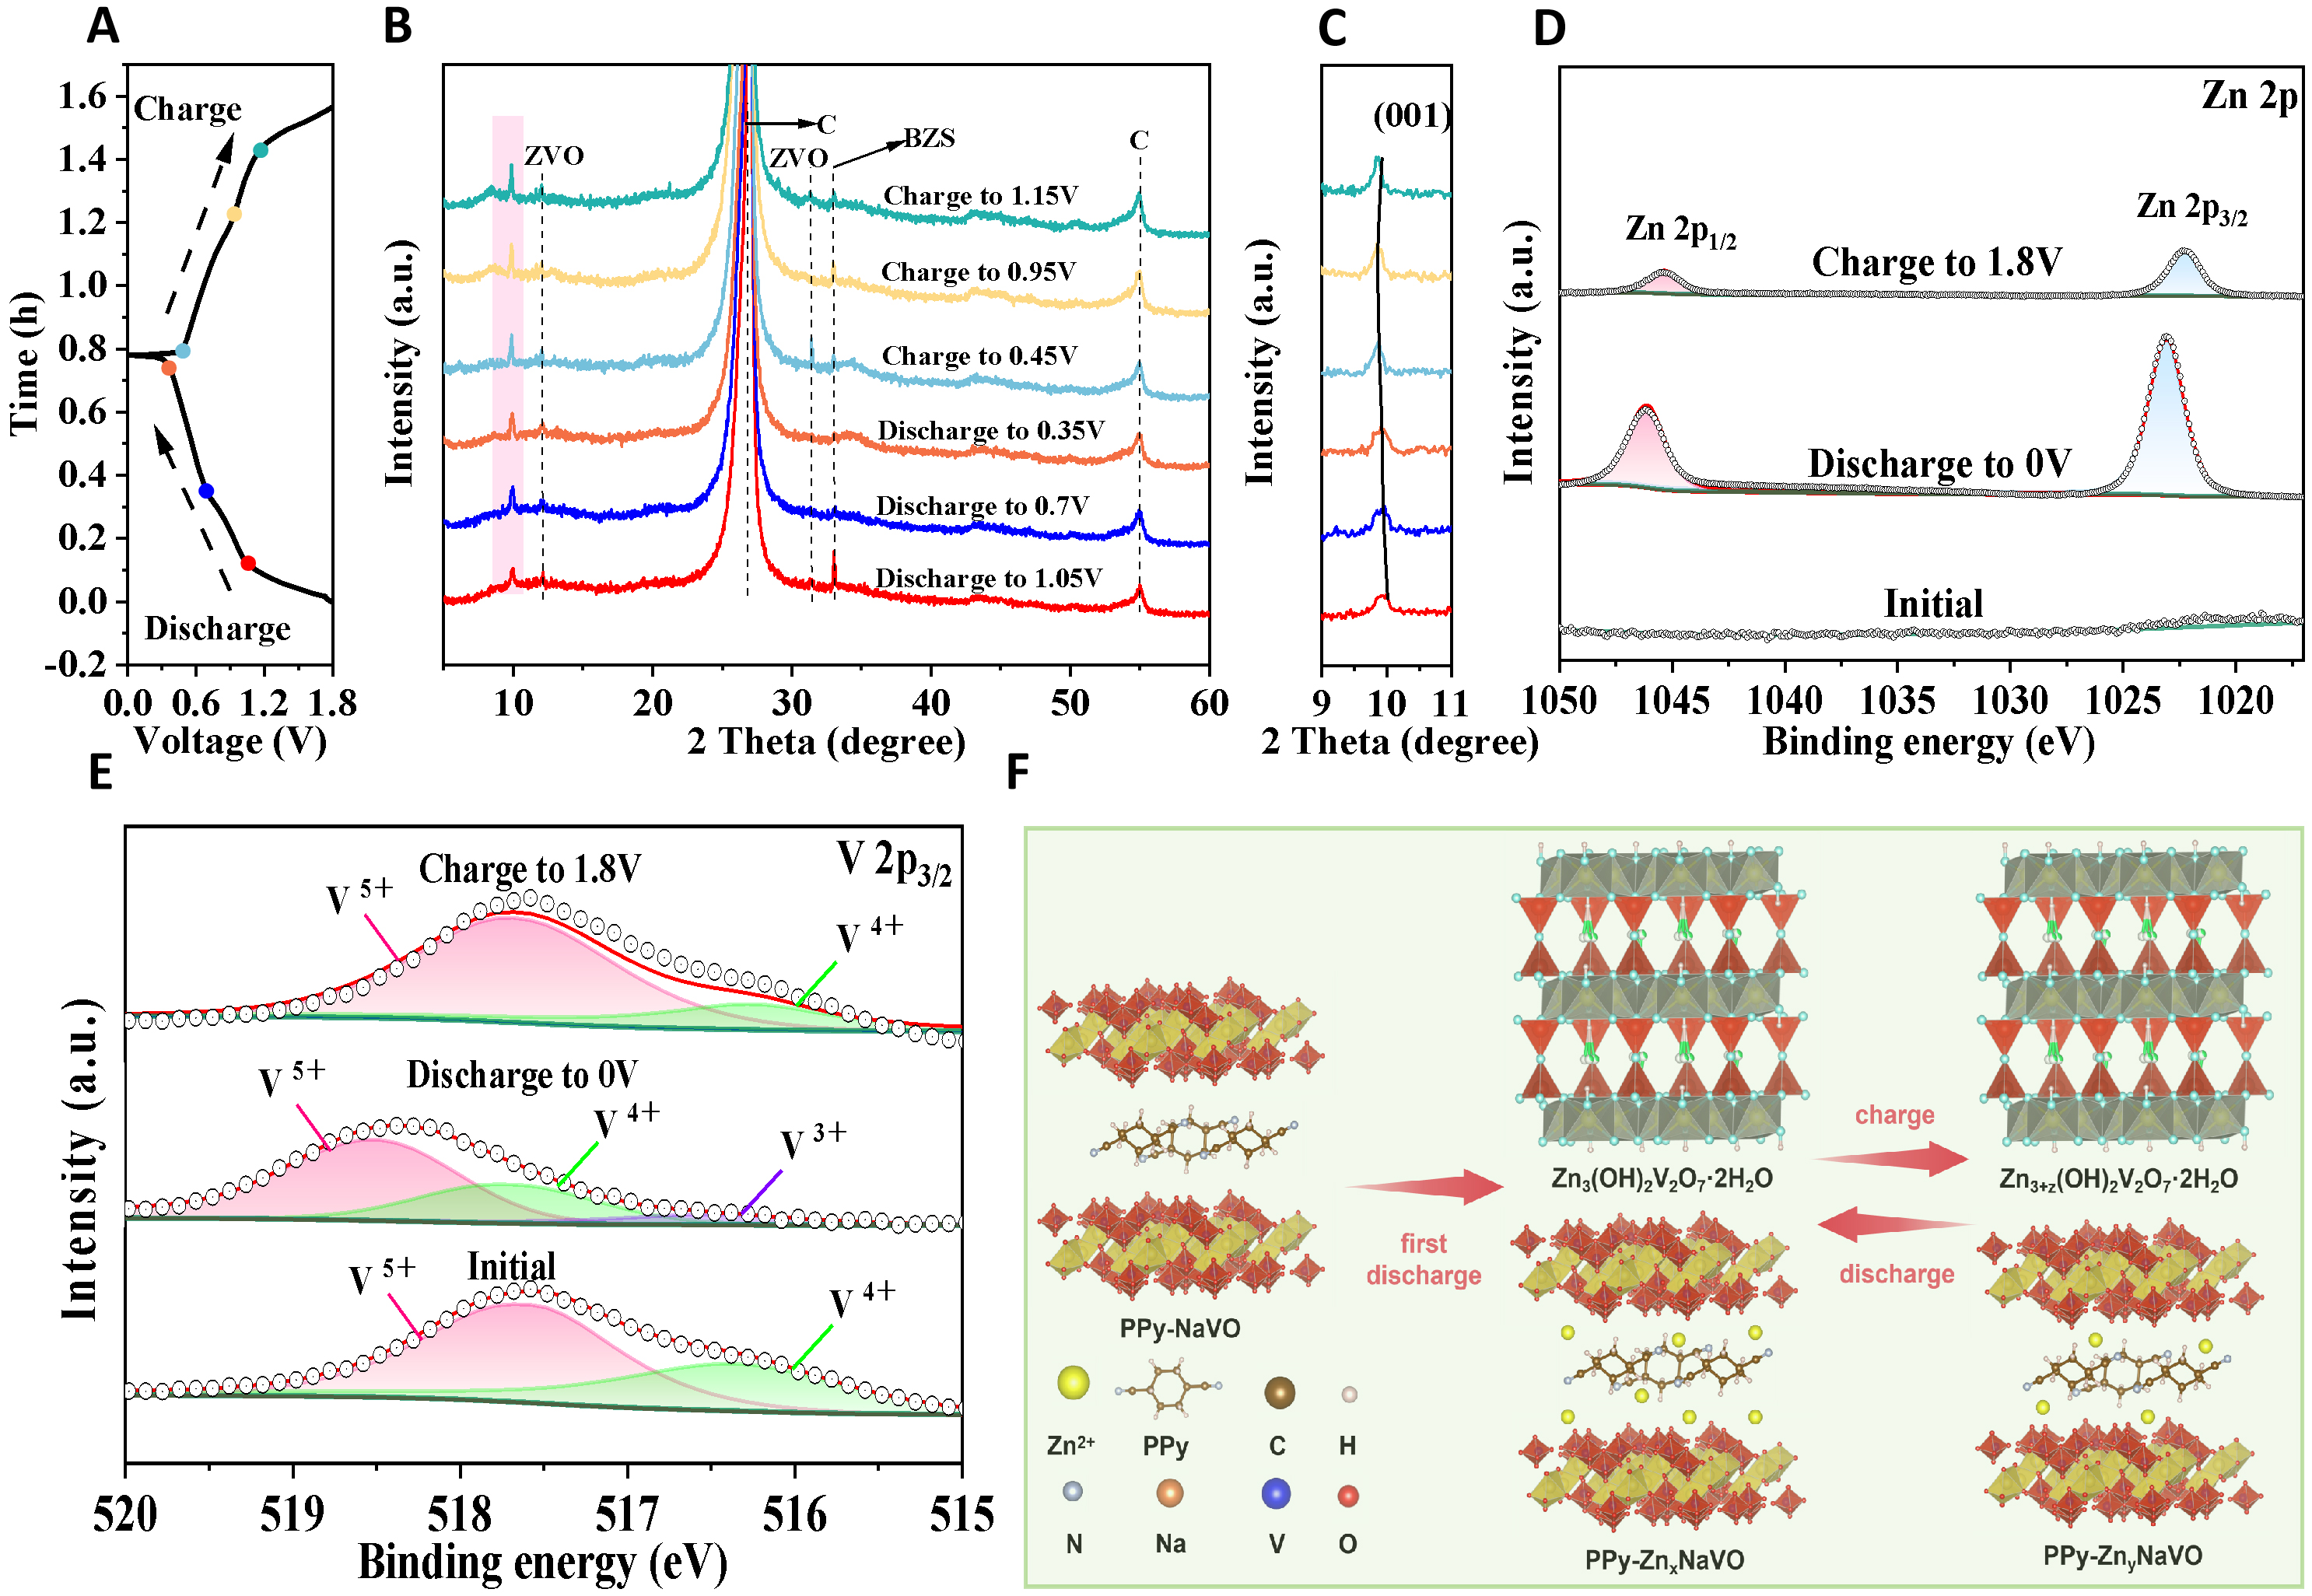
Figure 6. (A) Typical GCD profile of the electrode; Ex-situ analyses of XRD patterns at the selected points of (B) PPy-NaVO; (C) Enlarged view of the (001) diffraction peak; (D and E) Ex-situ XPS analyses; (F) Schematic illustration of the electrochemical storage process. GCD: Galvanostatic charge-discharge; XRD: X-ray diffraction; XPS: X-ray photoelectron spectroscopy; ZVO: Zn3(OH)2V2O7·2H2O; BZS: basic zinc salt; PPy-NaVO: polypyrrole-pillared Na2V6O16.
CONCLUSIONS
In conclusion, a PPy-NaVO composite was developed via a hydrothermal process and used as a cathode. The structural and electronic properties of the vanadate host are intrinsically optimized through polymer intercalation and the induction of a V4+/V5+ mixed-valence state. Structural analyses confirm that the intercalated PPy chains increase the interlayer distance from 0.79 nm to 0.89 nm. This expanded coordination environment, coupled with the formation of V4+ species, facilitates multi-electron redox reactions at the vanadium active centers. Ex-situ characterizations demonstrate that charge storage proceeds via a highly reversible Zn2+/H+ co-intercalation mechanism. During continuous cycling, the expanded interlayer spacing provides sufficient free volume to accommodate anisotropic lattice strain, while the continuous electronic pathways of the PPy network and the structural buffering framework help mitigate the structural collapse typically induced by the high charge density of Zn2+. This work provides an effective structural engineering method for building stable vanadium-based cathodes for aqueous energy storage.
DECLARATIONS
Acknowledgments
This work was partially supported by the Raman and BET tests provided by Scientific Compass (www.shiyanjia.com).
Authors’ contributions
Conceptualization, investigation, methodology, writing - original draft: Ying, J.; Mi, W.
Data curation, software, validation: Zhao, Y.; Liu, Y.; Li, J.
Formal analysis: Guo, C.; Zhang, S.
Resources, investigation, writing - review and editing: Dong, C.; Yin, B.; Ma, T.
Funding acquisition, project administration, supervision: Yin, B.; Ma, T.
Availability of data and materials
The original contributions presented in this study are included in the article/Supplementary Materials. Further inquiries can be directed to the corresponding author(s).
AI and AI-assisted tools statement
Not applicable.
Financial support and sponsorship
This work was supported by the Key Research Project of Department of Education of Liaoning Province (LJKZZ20220015), the Natural Science Foundation of Liaoning Province (2024-MS-007, 2024-MSLH-184), the Liaoning University Youth Research Fund project (LDZDJB2302), the Fundamental Research Funds for Public Universities in Liaoning (LJKLJ202425, LJ232410140033, LJ242510140014), Ma, T. acknowledges financial support from the Australian Research Council (ARC) through Future Fellowship (FT210100298), Discovery Project (DP220100603), Linkage Project (LP210200504, LP220100088, LP230200897), and Industrial Transformation Research Hub (IH240100009) schemes; the Australian Government through the Cooperative Research Centres Projects (CRCPXIII000077); the Australian Renewable Energy Agency (ARENA) as part of ARENA’s Transformative Research Accelerating Commercialisation Program (TM021); and the European Commission’s Australia-Spain Network for Innovation and Research Excellence (AuSpire).
Conflicts of interest
Zhang, S. is the Guest Editor of the special topic “Interfacial and Composite Material Design for Advanced Batteries, Photocatalysis, and Electrocatalysis” of the journal Energy Materials, but was not involved in any steps of editorial processing, notably including reviewer selection, manuscript handling, and decision-making, while the other authors have declared that they have no conflicts of interest.
Ethical approval and consent to participate
Not applicable.
Consent for publication
Not applicable.
Copyright
© The Author(s) 2026.
Supplementary Materials
REFERENCES
1. Jiang, T.; Shen, D.; Zhang, Z.; et al. Battery technologies for grid-scale energy storage. Nat. Rev. Clean. Technol. 2025, 1, 474-92.
2. Zhuang, C.; Zhang, S.; Yu, Z.; et al. Synergistic effect enables aqueous zinc-ion batteries to operate at high temperatures. Adv. Funct. Mater. 2025, 35, 2419351.
3. Zhou, Z.; Bian, J.; Yu, Z. Incremental cost analysis model of distribution network based on economic dispatch of distributed new-energy storage system. Clean. Energy. 2024, 8, 89-103.
4. Mishra, Y.; Chattaraj, A.; Aljabali, A. A.; El-tanani, M.; Tambuwala, M. M.; Mishra, V. Graphene oxide-lithium-ion batteries: inauguration of an era in energy storage technology. Clean. Energy. 2024, 8, 194-205.
5. Ngoy, K. R.; Lukong, V. T.; Yoro, K. O.; et al. Lithium-ion batteries and the future of sustainable energy: A comprehensive review. Renew. Sustain. Energy. Rev. 2025, 223, 115971.
6. Huang, Z.; Lai, J.; Kong, X.; et al. A solvent-anchored non-flammable electrolyte. Matter 2023, 6, 445-59.
7. Ye, Z.; Li, J.; Li, Z. Recent progress in nonflammable electrolytes and cell design for safe Li-ion batteries. J. Mater. Chem. A. 2023, 11, 15576-99.
8. Li, C.; He, P.; Zhang, T.; et al. Electron-driven-proton intercalation in δ-MnO2 to unlock fast-charging aqueous zinc-ion batteries. Adv. Energy. Mater. 2026, 16, e06142.
9. Tang, M.; Liu, Q.; Zou, X.; Zhang, B.; An, L. High-energy-density aqueous zinc-ion batteries: recent progress, design strategies, challenges, and perspectives. Adv. Mater. 2025, 37, e2501361.
10. Lv, W.; Tan, Y.; Guo, C.; et al. Triple regulation of water molecules behavior to realize high stability and broad temperature tolerance in aqueous zinc metal batteries via a novel cost-effective eutectic electrolyte. Adv. Energy. Mater. 2025, 15, 2403689.
11. Bag, S.; Thakur, V. S.; Bhadra, A.; Banerjee, S.; Kundu, D.; Raj, C. R. Vacancy-engineered vanadium cathode for fast-charging aqueous zinc batteries of ultra-long cycle Life. Adv. Funct. Mater. 2026, 36, e15788.
12. Zhang, X.; Xu, X.; Liu, S.; et al. Isometric epilayer strategy via lattice matching enabling polycrystalline vanadium oxide for stable and ultrafast zinc-ion storage. J. Energy. Chem. 2026, 120, 561-71.
13. Hu, X.; Gao, S.; Lin, T.; et al. Iodine-doped sodium vanadate cathode for improved Zn ion diffusion kinetics. Adv. Mater. 2025, 37, e2416714.
14. Gan, L.; Liu, F.; Yuan, X.; Fu, L.; Wu, Y. Alumina modified sodium vanadate cathode for aqueous zinc-ion batteries. Front. Energy. 2023, 17, 775-81.
15. Guo, X.; Fang, G.; Zhang, W.; et al. Mechanistic insights of Zn2+ storage in sodium vanadates. Adv. Energy. Mater. 2018, 8, 1801819.
16. Yang, J.; Zhang, Y.; Ge, Y.; et al. Interlayer engineering of layered materials for efficient ion separation and storage. Adv. Mater. 2024, 36, e2311141.
17. Zhou, X.; Yang, C.; Liu, X.; et al. Inhibiting inter-layer gliding in transition metal layered oxides through interphase engineering for sodium-ion batteries. Nat. Commun. 2025, 16, 6691.
18. Guo, J.; Meng, S.; Yan, X.; et al. Interlayer-engineered ammonium vanadate cathodes via Na+/PEG Co-intercalation for fast and stable Zn-ion storage. Chem. Eng. J. 2026, 531, 173885.
19. Han, J.; Gao, S.; Sun, Z.; Yang, Z.; Liu, X.; Wang, C. Layered structure modification of sodium vanadate through Ca/F co-doping for enhanced energy storage performance. Adv. Energy. Mater. 2024, 14, 2401481.
20. Li, X.; Wang, L.; Fu, Y.; Dang, H.; Wang, D.; Ran, F. Optimization strategies toward advanced aqueous zinc-ion batteries: from facing key issues to viable solutions. Nano. Energy. 2023, 116, 108858.
21. Chen, K.; Zong, Q.; Liu, X.; et al. Surface- and interlayer-modified ammonium vanadate cathode for high-performance aqueous Zn-ion batteries. J. Colloid. Interface. Sci. 2025, 693, 137587.
22. He, Q.; Wang, Y.; Wu, Q.; et al. Phase-transition intervention engineered amorphous vanadium oxide for high-performance aqueous zinc-ion batteries. Angew. Chem. Int. Ed. 2025, 64, e17305.
23. Liu, Y.; Wang, T.; Sun, Y.; et al. Fast and efficient in-situ construction of low crystalline PEDOT-intercalated V2O5 nanosheets for high-performance zinc-ion battery. Chem. Eng. J. 2024, 484, 149501.
24. Yu, N.; Zeng, Q.; Hu, L.; Guo, K.; Wang, L. Low-strain cathode via electrochemical conversion for high-power and durable aqueous zinc-ion batteries. ACS. Nano. 2026, 20, 1278-89.
25. Wang, Y.; Chen, X.; Liu, J. H.; et al. Dual-functional nanoengineering via molecular pillaring and conductive hybridization for high-performance aqueous zinc-ion batteries. Chem. Sci. 2026, 17, 5115-24.
26. Cao, S.; Lu, Y.; Tang, Y.; et al. Constructing ion-transport blockchain by polypyrrole to link CoTi-ZIF-9 derived carbon materials for high-performance seawater desalination. J. Colloid. Interface. Sci. 2024, 654, 466-75.
27. Feng, Z.; Sun, J.; Liu, Y.; et al. Polypyrrole-intercalation tuning lamellar structure of V2O5·nH2O boosts fast zinc-ion kinetics for aqueous zinc-ion battery. J. Power. Sources. 2022, 536, 231489.
28. Gao, Q.; Song, C.; Zhang, W.; et al. Ordered organic-inorganic dual-layer synergistic interfaces on zinc anodes enable high-performance zinc metal batteries. Adv. Funct. Mater. 2026, 36, e76812.
29. Guo, D.; Fan, Y.; Yang, Q.; et al. Oxygen-deficient organic/inorganic VOx-PPy cathode composites: enabling prolonged lifespan of aqueous zinc-ion batteries. Chem. Eng. J. 2025, 507, 160745.
30. Wang, Z.; Wang, K.; Nie, K.; et al. Three-dimensional through-localized amorphization engineering of crystalline/amorphous heterophase ammonium vanadate nanobelts for advanced aqueous zinc-ion batteries. Adv. Funct. Mater. 2026, 36, e15102.
31. Zhang, D.; Luo, X.; Limphirat, W.; Qin, J.; Yang, X.; Cao, J. Synergistic d-π coupling and interfacial catalysis enable electron-ion dual kinetics in Ah-level zinc-ion batteries. Adv. Funct. Mater. 2026, 36, e76770.
32. Qi, Y.; Jin, X.; Xu, L.; Ren, X.; Xia, Y. Ultra-high mass-loading V5+-VO2@PPy cathode for aqueous zinc-ion battery. J. Alloys. Compd. 2023, 967, 171750.
33. Liu, C.; Meng, Z.; Cai, Y.; et al. In situ aniline polymerization during NH4V4O10 lattice formation achieves a high-performance hybrid cathode for aqueous zinc-ion batteries. Chem. Commun. 2024, 60, 14838-41.
34. Lascelles, S.; Armes, S.; Zhdan, P.; et al. Surface characterization of micrometre-sized, polypyrrole-coated polystyrene latexes: verification of a ‘core–shell’ morphology. J. Mater. Chem. 1997, 7, 1349-55.
35. Huang, H.; Xia, X.; Yun, J.; et al. Interfacial engineering of hydrated vanadate to promote the fast and highly reversible H+/Zn2+ co-insertion processes for high-performance aqueous rechargeable batteries. Energy. Storage. Mater. 2022, 52, 473-84.
36. Zhou, H.; Xu, X.; Liang, P.; et al. Low-temperature-resilient and high-rate polypyrrole-vanadium oxide carbon fiber structural supercapacitors with superior mechanical-electrochemical synergy. Adv. Energy. Mater. 2026, 16, e70890.
37. Yang, Y.; Tang, Y.; Liang, S.; et al. Transition metal ion-preintercalated V2O5 as high-performance aqueous zinc-ion battery cathode with broad temperature adaptability. Nano. Energy. 2019, 61, 617-25.
38. Liang, X.; Yang, Y.; Di, W.; et al. Three-dimensional mixed-valence polyoxovanadates@polyaniline architecture towards high-performance rechargeable aqueous zinc-ion batteries cathode. Chem. Eng. J. 2024, 495, 153255.
39. Wang, T.; Li, S.; Weng, X.; et al. Ultrafast 3D hybrid-ion transport in porous V2O5 cathodes for superior-rate rechargeable aqueous zinc batteries. Adv. Energy. Mater. 2023, 13, 2204358.
40. Zhou, T.; Du, X.; Gao, G. Revealing the role of calcium ion intercalation of hydrated vanadium oxides for aqueous zinc-ion batteries. J. Energy. Chem. 2024, 95, 9-19.
Cite This Article
 Open Access
How to Cite
Ying, J.; Mi, W.; Zhao, Y.; Zhang, S.; Liu, Y.; Li, J.; Guo, C.; Dong, C.; Ma, T.; Yin, B. Hierarchical polypyrrole-pillared Na2V6O16 architectures enabling high-rate and stable aqueous zinc storage. Energy Mater. 2026, 6, 600147. https://dx.doi.org/10.20517/energymater.2026.240
Download Citation
If you have the appropriate software installed, you can download article citation data to the citation manager of your choice. Simply select your manager software from the list below and click on download.
Export Citation File
Type of Import
Tips on Downloading Citation
Citation Manager File Format
Type of Import
Direct Import: When the Direct Import option is selected (the default state), a dialogue box will give you the option to Save or Open the downloaded citation data. Choosing Open will either launch your citation manager or give you a choice of applications with which to use the metadata. The Save option saves the file locally for later use.
Indirect Import: When the Indirect Import option is selected, the metadata is displayed and may be copied and pasted as needed.
About This Article
Special Topic
Copyright

Data & Comments
Data
0






Comments
Comments must be written in English. Spam, offensive content, impersonation, and private information will not be permitted. If any comment is reported and identified as inappropriate content by OAE staff, the comment will be removed without notice. If you have any queries or need any help, please contact us at [email protected].